
ISO 13485 quality control checklist items that get missed first
Publication Date
author
In ISO 13485 audits, the first failures rarely come from obvious gaps—they come from overlooked details inside the iso 13485 quality control checklist, especially where documentation, traceability, validation, and process controls intersect. For teams managing micro machining tolerance limits, edm surface integrity analysis, or medical grade peek sterilization test requirements, missing these early checklist items can quickly escalate from minor nonconformities to supplier-risk and compliance delays.
For engineering teams, operators, procurement managers, and executive decision-makers, the challenge is not simply “having a quality system.” The challenge is whether the system can withstand scrutiny at the exact point where design intent, manufacturing execution, and evidence records meet. In precision machining and regulated component supply, the items missed first are usually small on paper but high-impact in an audit.
This article breaks down the checklist items that tend to fail early, why they matter in real hard-tech environments, and how to turn them into measurable controls. The focus is practical: documentation discipline, process validation boundaries, traceability depth, supplier oversight, and inspection evidence that procurement and technical teams can actually use when qualifying or comparing manufacturers.
Where the First ISO 13485 Quality Control Checklist Gaps Usually Appear
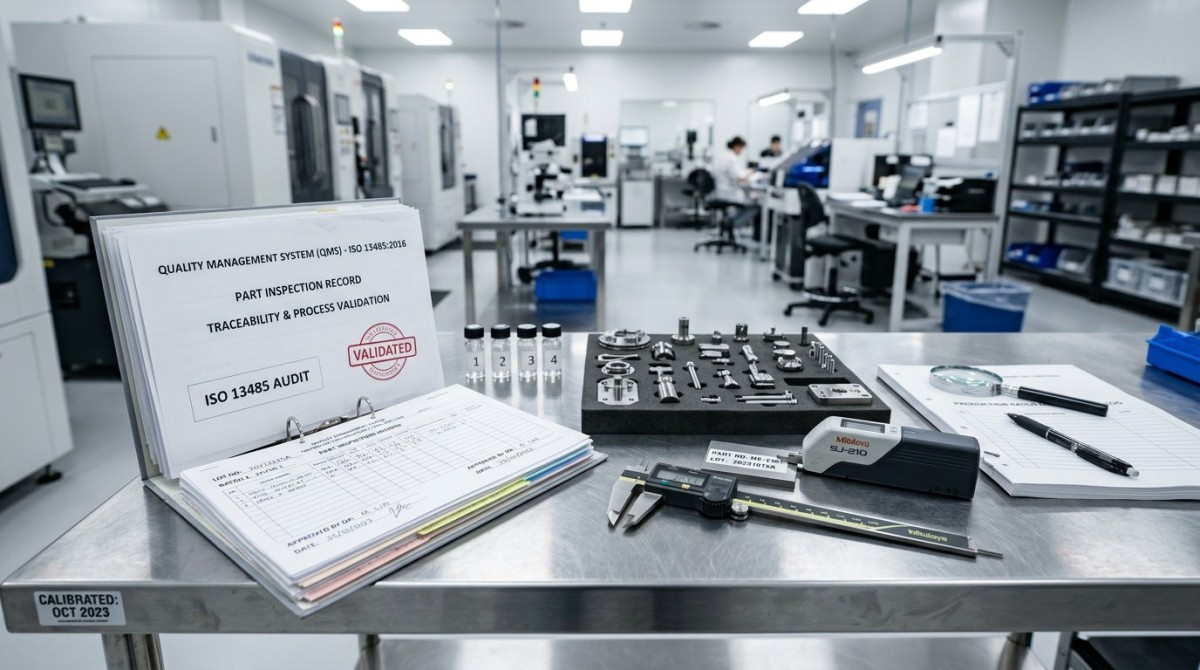
The first missed items in an iso 13485 quality control checklist are rarely dramatic. Auditors often identify issues in 4 early zones: uncontrolled document changes, incomplete device history records, vague acceptance criteria, and unclosed validation assumptions. In a machining-heavy medical supply chain, these weak points surface before larger system failures because they are easy to sample and difficult to defend without objective evidence.
A common example is the mismatch between drawing revision, router version, and inspection plan. If a CNC process sheet references Rev. B, but the final inspection report uses Rev. C tolerances, the problem is not just clerical. It creates uncertainty over which specification was actually manufactured. In regulated production, even a ±0.005 mm difference can trigger a deeper review when critical-to-quality features are involved.
Another frequent gap appears in validation language. Teams may state that a cleaning, sterilization preparation, or EDM finishing process is “validated,” yet cannot define the tested limits. Auditors usually ask for the upper and lower boundaries: load size, cycle count, exposure time, material lot variation, and requalification trigger. If those limits are missing, the validation is often viewed as incomplete rather than robust.
Operators also encounter early findings when in-process checks are performed but not linked to reaction plans. Recording a surface roughness value of Ra 0.4 µm has little compliance value if the work instruction does not state what happens at Ra 0.45 µm, who approves disposition, and how recurrence is prevented within the next 24 to 72 hours of production.
For procurement and supplier-quality teams, these early misses matter because they indicate systemic maturity. A supplier can present modern 5-axis equipment, cleanroom capability, and fast lead times of 2 to 4 weeks, yet still fail on basic quality evidence. That mismatch is exactly where qualification cycles become longer and sourcing risk increases.
The difference between visible compliance and usable control
Visible compliance means the procedures exist. Usable control means the records, revision links, inspection criteria, and deviation responses all align under sampling. In ISO 13485 environments, auditors normally test usability, not just document presence. If a system cannot show who changed a record, when it changed, and what downstream documents were updated within a defined interval such as 48 hours, it is vulnerable.
High-risk early-miss signals
- Inspection forms without revision status, approval signature, or acceptance limit source.
- Traceability records that stop at batch level when serial-level identification is required.
- Validated process claims without documented worst-case conditions or revalidation criteria.
- Nonconformance reports opened within 7 days but lacking closure evidence or effectiveness review.
Documentation and Traceability Items That Auditors Check First
Documentation and traceability are often the fastest route to an audit finding because they reveal whether the quality system works across departments. In precision manufacturing, the issue is not only whether a part passed final inspection. The issue is whether the organization can reconstruct the full manufacturing story: material source, machine program revision, operator authorization, inspection method, and disposition history for each lot or serial number.
This is especially important when dealing with specialty polymers, implant-adjacent components, micro-machined housings, or EDM-finished geometries. If a team cannot connect raw material certificates to heat number, incoming acceptance, machining traveler, post-process cleaning, and final release record, the traceability chain is incomplete. Auditors often sample backward from shipment to receipt and forward from receipt to shipment in 2 separate trace exercises.
One overlooked area is metadata consistency. A record may be technically complete but still weak if part number formatting, lot code logic, and revision naming differ between ERP, MES, and paper travelers. That inconsistency creates avoidable confusion during supplier transfer, CAPA review, or complaint investigation. In practice, one formatting mismatch can cost several hours in an audit room and much more during a field issue review.
Another missed item is retention logic. ISO 13485-driven environments usually define retention periods based on product type, contractual requirement, and regulatory context. If the organization keeps calibration records for 1 year but retains product history for 5 years, there may be no way to confirm measurement validity for an older batch. That gap becomes serious when complaint trends emerge 18 to 36 months after shipment.
Checklist items that deserve immediate review
The table below highlights documentation and traceability points that are commonly missed first, along with the practical risk each gap creates for engineering, supplier quality, and procurement teams.
The key takeaway is that traceability failure is usually cumulative rather than singular. A missing operator signature may be manageable. A missing signature combined with no gauge identification, unclear lot coding, and mixed document revisions becomes an audit pattern. That pattern raises questions about control culture, not just one isolated mistake.
Practical control steps
- Standardize part number, lot code, and revision syntax across ERP, MES, and quality records.
- Require each inspection record to include gauge ID, method reference, and spec revision.
- Run monthly backward-and-forward trace drills on at least 2 sampled production lots.
- Align retention periods so that calibration, process, and product records remain reviewable together.
Validation and Process Control Weak Points in Precision Manufacturing
Validation is one of the most misunderstood parts of the iso 13485 quality control checklist. Teams often assume that if a process is documented and repeated, it is effectively validated. In reality, auditors want to see objective proof that the process performs within defined limits over time, across realistic production conditions, and with known inputs. This matters for CNC machining, EDM, cleaning, packaging preparation, and sterilization-related preconditioning steps.
Take EDM surface integrity analysis as an example. A supplier may confirm dimensional accuracy, but not document the effect of recast layer thickness, microcracking risk, or downstream cleaning adequacy on the final part. If the process window is not tied to measurable outputs, the validation remains too narrow. For critical applications, organizations often define limits such as spark energy range, electrode wear threshold, and post-EDM surface roughness acceptance before release.
The same issue appears in medical grade PEEK processing. Material identity may be controlled, but teams sometimes fail to document the sterilization test interface: temperature exposure range, cycle count tolerance, packaging interaction, and dimensional stability after repeated conditioning. Even when the organization does not perform sterilization itself, its quality control checklist should define how material performance evidence is reviewed and retained from qualified partners.
Process control also breaks down when reaction plans are generic. If a spindle load trend exceeds an internal alert limit by 10%, does production stop immediately, shift to 100% inspection, or continue under deviation? Without that decision tree, control is informal. In regulated production, informal control is difficult to defend because repeatability depends too much on individual judgment rather than approved criteria.
For procurement teams assessing a supplier, validation maturity often predicts long-term consistency better than machine count or marketing claims. A smaller supplier with 3 validated critical processes and defined requalification intervals of 12 months may be lower risk than a larger one with advanced equipment but weak process boundaries and inconsistent evidence retention.
What robust validation records usually include
The following comparison shows the difference between superficial process claims and auditable process validation controls in hard-tech medical manufacturing environments.
The main pattern is clear: strong validation records define boundaries, evidence, and response paths. Weak records rely on general statements. During supplier qualification, buyers should ask to see not only the procedure, but also one executed validation file, one recent revalidation decision, and one deviation example from the last 6 to 12 months.
Four process-control questions buyers should ask
- What are the upper and lower validated operating limits for this process?
- What event triggers revalidation: time, tooling change, software update, or material source change?
- How often are in-process checks performed: every piece, every 30 pieces, or per defined time interval?
- What documented reaction plan applies when results drift but remain technically within tolerance?
Supplier Control, Purchasing Oversight, and Audit-Ready Acceptance Criteria
Purchasing controls are often underestimated in ISO 13485 audits, yet supplier quality is one of the fastest ways for a nonconformity to expand beyond one site. If a company buys precision-machined parts, sterilization support services, specialty polymers, or outsourced finishing, its iso 13485 quality control checklist should define how suppliers are approved, monitored, and re-evaluated. A purchase order alone is not a sufficient control mechanism.
A recurring issue is that supplier approval is based on commercial history instead of technical risk. For low-risk hardware, that may be manageable. For medical-grade precision parts, however, the evaluation usually needs at least 3 dimensions: process capability, documentation discipline, and change-notification reliability. If a supplier changes a sub-tier raw material source or inspection method without notice, the customer may inherit compliance risk before realizing it.
Acceptance criteria also deserve closer attention. Many organizations define what “pass” means on the drawing but fail to define how acceptance is demonstrated in receiving inspection or supplier scorecards. For example, should incoming lots be checked by ANSI-style sampling, 100% dimensional review of critical features, or certificate verification plus periodic dock audit? If that logic is not defined, incoming quality becomes inconsistent from one inspector or buyer to another.
From a procurement perspective, the most effective supplier controls are measurable and repeatable. Late delivery rate, documentation error rate, response time to NCR, on-time change notification, and closure time for corrective actions all matter. A supplier with a 98% on-time delivery rate but repeated document mismatches may still be high risk in regulated medical production.
For executive teams, this is not only about compliance. Strong supplier control reduces qualification cycles, shortens dispute resolution, and improves launch predictability when new parts move from prototype to production over a 3-stage ramp: first article, pilot lot, and serial release.
Supplier oversight metrics worth adding to the checklist
The table below translates supplier control into practical review points that both procurement and quality teams can use during onboarding and quarterly business review.
These controls help separate strong suppliers from merely responsive ones. In hard-tech sourcing, responsiveness is useful, but evidence quality is what protects long-term production. Teams that combine supplier metrics with technical reviews usually make faster and more defensible sourcing decisions.
Minimum purchasing-control checks
- Define supplier approval by risk category rather than by spend alone.
- Attach drawing revision, acceptance criteria, and traceability expectations directly to purchasing documentation.
- Review at least quarterly whether supplier changes, NCR closure, and incoming inspection trends remain aligned.
- Require escalation thresholds when repeat defects exceed a set frequency, such as 2 lots in 90 days.
How to Build a Practical Review Routine Before the Audit Starts
The most effective way to avoid early misses in an iso 13485 quality control checklist is to convert audit clauses into recurring operating reviews. Instead of waiting for an annual external audit, teams should review evidence monthly, by process family, and by risk. This approach is especially useful for manufacturers handling high-mix, low-volume medical parts where documentation variation can build quickly across many small jobs.
A practical routine begins with 5 review buckets: document control, traceability, process validation, equipment and calibration, and supplier performance. Each bucket should include one sample-based check and one trend-based check. For example, sample 3 recent device history records and trend 6 months of NCR closure times. This method balances detail with management visibility.
Cross-functional review is equally important. Operators know where real workarounds happen. Quality engineers know where records fail. Procurement sees where supplier communication breaks. Decision-makers need this combined view because many early findings are interface failures rather than single-department failures. A 30- to 60-minute monthly review can reveal issues that would otherwise remain hidden until audit day.
For teams working with micro machining tolerance limits or specialized materials, one valuable technique is the “evidence walk.” Start from a shipped lot and walk backward through final inspection, in-process checks, setup approval, material receipt, and approved supplier status. Then repeat the walk forward from material receipt to shipment. If either path breaks, the checklist needs correction before the auditor performs the same exercise.
Organizations that treat audit readiness as an operational habit usually reduce both compliance friction and sourcing uncertainty. They can compare suppliers more clearly, support engineering change decisions faster, and shorten qualification discussions because the evidence package already exists in usable form.
A simple pre-audit operating rhythm
- Weekly: review 1 controlled document change and 1 nonconformance closure for completeness.
- Monthly: sample 2 to 3 production lots for traceability depth and inspection record integrity.
- Quarterly: review supplier scorecards, process changes, and overdue CAPA actions.
- Every 6 to 12 months: confirm validation status, requalification triggers, and training effectiveness for critical processes.
FAQ: common decision-stage questions
How detailed should an ISO 13485 checklist be for a precision machining supplier?
It should go beyond a clause-level list. At minimum, include document revision control, lot or serial traceability, validation boundaries for special processes, gauge linkage, supplier-change notification rules, and reaction plans for out-of-trend results. In many cases, 20 to 40 operational checkpoints are more useful than a short generic checklist.
What records are most useful during supplier qualification?
Ask for one recent device history record, one executed validation package, one nonconformance with corrective action, one calibration sample, and one traceability demonstration from incoming material to final shipment. These 5 record types often reveal more than a broad slide presentation.
How often should supplier performance be re-evaluated?
For critical suppliers, quarterly review is a practical baseline. If defect recurrence, documentation errors, or change-control problems increase, a monthly review may be necessary until the trend stabilizes for at least 90 days.
The fastest audit failures do not come from the most complicated requirements. They come from the first missing links between specification, execution, and evidence. In any iso 13485 quality control checklist, the items most often missed first are document revision accuracy, traceability continuity, validation boundaries, supplier-control discipline, and reaction plans tied to measurable acceptance criteria.
For organizations sourcing or producing precision medical components, these controls are not paperwork overhead. They are the basis for reliable qualification, faster engineering reviews, and lower supplier risk. TechStat Vanguard focuses on this exact decision layer: separating technical substance from marketing language so buyers and engineering teams can judge capability through measurable control, not broad claims.
If your team is reviewing supplier readiness, refining a medical manufacturing checklist, or comparing process-control maturity across vendors, now is the right time to tighten the evidence chain before the next audit or sourcing decision. Contact us to discuss a tailored evaluation framework, request a technical review structure, or explore more hard-tech quality and benchmarking insights.
Search News
Hot Articles
- Carbon Fiber StructuresHow to Specify Aerospace Carbon Fiber Assemblies for Lightweight, Load-Bearing PartsJul 11, 2026
- Machine VisionXiaomi, Alibaba DAMO Academy Drive Open-Source Robotics Model Ecosystem, Lowering Algorithm Development Barriers for Overseas SMEs Adopting Chinese Robotics SolutionsMar 30, 2026
- Cobots & ArmsShanxi 'AI+' Conference Showcases Embodied Intelligence with Multinational Buyer Engagement: Key Industry ImplicationsMar 30, 2026
Popular Tags
Recommended News
 Jul 11, 2026How to Specify Aerospace Carbon Fiber Assemblies for Lightweight, Load-Bearing PartsAerospace carbon fiber assemblies: learn how to specify laminate design, load data, joining methods, and inspection controls for lighter, reliable load-bearing aerospace parts.Elena Rostova (UAV Systems Researcher)
 Mar 30, 2026Xiaomi, Alibaba DAMO Academy Drive Open-Source Robotics Model Ecosystem, Lowering Algorithm Development Barriers for Overseas SMEs Adopting Chinese Robotics SolutionsXiaomi & Alibaba DAMO Academy's open-source robotics models lower algorithm barriers for global SMEs. Boost efficiency in manufacturing, logistics & smart retail with ROS 2 compatible solutions.TSV Data Lab
 Mar 30, 2026Shanxi 'AI+' Conference Showcases Embodied Intelligence with Multinational Buyer Engagement: Key Industry ImplicationsShanxi 'AI+' Conference highlights embodied intelligence with Tesla, Ubtech & Jiantao showcasing multimodal robotics. 40+ global buyers validated tech on-site, creating new B2B channels for industrial automation & AI solutions.Chen Wei (Automation Lead Engineer)
 Mar 30, 2026Zhongguancun Unveils 6G Integrated Sensing-Computing-Intelligence Platform to Empower Embodied Robots for Global Industrial ApplicationsZhongguancun's new 6G platform revolutionizes industrial robotics with <10ms latency, empowering global automotive, logistics & port automation. Discover how this tech boosts China's smart manufacturing exports.TSV Data Lab
 Mar 30, 2026Mech-Mind Tianjin Embodied Intelligence R&D and Production Base Goes Live, Cutting Delivery Time by 20%–30% for Global Customized AMR+Vision SolutionsMech-Mind's Tianjin AMR+Vision production base cuts delivery time by 20%-30% for欧美 & Latin American markets. Discover how this PLd-certified facility accelerates customized robotics solutions for smart manufacturing and logistics.Chen Wei (Automation Lead Engineer)
 Mar 30, 2026Boao Forum Reveals 'Co-evolution of Big and Small Brains' Trend in Humanoid Robots, China's Embodied Intelligence Accelerates Adaptation to Overseas Manufacturing Service ScenariosDiscover how China's humanoid robots, featuring 'Co-evolution of Big and Small Brains,' are revolutionizing global manufacturing. Learn about Embodied Tian Gong 3.0's ISO compliance and overseas deployment at Boao Forum 2026.Chen Wei (Automation Lead Engineer)
 Mar 30, 2026China Railway Industrial Secures Patent for Embodied Robotics in Material Handling, Boosting High-Precision Delivery Capabilities for Overseas Infrastructure and Manufacturing ProjectsChina Railway Industrial's new patent for embodied robotics in material handling boosts high-precision delivery for global infrastructure & manufacturing. Discover how this innovation enhances logistics accuracy in automotive, rail transit & EPC projects.Chen Wei (Automation Lead Engineer)
 Oct 27, 2026GXO and Agility Robotics Expand Partnership to Accelerate US Humanoid Robot DeploymentGXO and Agility Robotics plan to expand US humanoid robot deployment with Digit 4 in early 2027. Explore warehouse automation plans, service terms and future Digit 5 upgrades.Chen Wei (Automation Lead Engineer)
 Oct 27, 2026Robotic Automation Costs: Budgeting for Equipment, Integration, and MaintenancePlan your robotics automation cost with confidence. Explore equipment, integration, and maintenance expenses, compare quotes, and build a realistic budget for better ROI.Chen Wei (Automation Lead Engineer)
 Oct 26, 2026AMAXiMAL Releases Open-Source TopMod 3D v3 With Claude AIExplore AMAXiMAL’s free, open-source TopMod 3D v3 with Claude AI assistance, topology-based modeling and plywood fabrication tools that connect complex designs to physical builds.Dr. Marcus Vance
 Oct 26, 2026Choosing a SCADA System Integrator Company: PLC Compatibility and Long-Term SupportChoose a scada system integrator company with proven PLC compatibility and reliable long-term support. Discover what to test, compare, and require before investing.Victor Lin (Chief Software Architect)
 Oct 25, 2026Hesai Leads Global ADAS Main Lidar Shipments in Yole’s 2026 ReportHesai led global ADAS main lidar shipments with a 43% share in 2025, according to Yole’s 2026 report. Explore what this ranking reveals about its automotive market position.TSV Data Lab
 Oct 25, 2026Choosing an Aerospace Materials Manufacturer: Quality and Traceability ChecksChoosing an aerospace materials manufacturer? Learn how to verify certifications, test reports, and lot traceability to qualify suppliers confidently and reduce costly sourcing risks.Dr. Marcus Vance
 Oct 24, 2026Guotai Haitong: Global Entry-Level 3D Printer Shipments Rose 39% in Q1 2026Entry-level 3D printer shipments rose 39% in Q1 2026, Guotai Haitong reports. Explore how AI, multicolor printing and print farms are shaping market growth.Dr. Marcus Vance
 Oct 24, 2026Choosing an Industrial Vision Inspection Supplier: Acceptance Tests That MatterChoose an industrial vision inspection supplier with confidence. Discover acceptance tests for defect detection, false rejects, production speed, and reliable part rejection before you buy.TSV Data Lab
 Oct 23, 2026US Navy Raises Gecko Robotics Contract Ceiling to $300M for Robotic Fleet MaintenanceUS Navy raises Gecko Robotics’ contract ceiling to $300M for Pacific Fleet robotic inspections. Explore potential opportunities for maintenance suppliers and key procurement considerations.Chen Wei (Automation Lead Engineer)
 Oct 23, 2026Choosing Industrial Sensing Systems for Global Plants: Protocol CompatibilityCompare industrial sensing systems international buyers can trust. Explore protocol compatibility, reduce integration risks, and select proven configurations for reliable global plant deployments.TSV Data Lab
 Oct 22, 2026KUKA Showcases Physical AI and iiQKA.AI Vision at NVIDIA GTC BerlinExplore KUKA’s Physical AI showcase at NVIDIA GTC Berlin and discover how iiQKA.AI Vision aims to simplify robotic vision integration and improve 2D and 3D scene recognition.TSV Data Lab
 Oct 22, 2026Robotic Machining of Advanced Composites: What Tolerances Are Achievable?Explore advanced materials machining robotics: discover what drives achievable composite tolerances and how to validate accuracy before investing in your next machining system.Dr. Marcus Vance
 Oct 21, 2026China’s Industrial Robot Exports Near RMB 10 Billion in First Nine Months of 2026China’s industrial robot exports neared RMB 10 billion in January–September 2026, with value rising faster than volume. Explore the data and what it could mean for suppliers.Chen Wei (Automation Lead Engineer)
 Oct 21, 2026Low-Latency Industrial Data Processing: Pricing and Key Cost DriversExplore industrial data latency price drivers, from edge hardware to integration and support. Learn how to compare quotes, verify performance, and avoid unnecessary costs.TSV Data Lab
 Oct 20, 2026KUKA Launches iiQKA.AI Vision to Simplify Industrial Image ProcessingKUKA iiQKA.AI Vision simplifies industrial image processing with controller-based 2D and 3D vision. Explore drag-and-drop setup for vision-guided automation.TSV Data Lab
 Oct 20, 2026Smart Industrial Edge Devices: Key Criteria for Legacy PLC IntegrationChoose smart industrial edge devices for legacy PLC integration. Discover key criteria for compatibility, data quality, security, and reliable collection without disrupting control.TSV Data Lab
 Oct 19, 2026Choosing a Radar Sensor by Application: Range, Resolution, and Field of ViewSimplify radar sensor application based product selection. Compare range, resolution, and field of view to avoid blind spots and choose with confidence. Explore the guide.TSV Data Lab























